China Best Medical Manufacturing What Standards Apply?

Time:2026-10-29 Author:Aria
0%

China’s medical manufacturing sector produces everything from molded diagnostic housings to powered surgical equipment. Each product brings different risks, processes, and market requirements. The question “what standards apply to medical manufacturing” has no single answer. It depends on the device, its intended use, and where it will be sold.

ISO 13485 is widely used for medical-device quality management, while ISO 14971 addresses risk management. Electrical equipment may need evaluation against relevant IEC 60601 requirements. Materials that contact patients may require biological-safety assessment, often guided by the ISO 10993 series. These standards do not apply identically to every product. China’s NMPA requirements matter for devices placed on the Chinese market; other destinations may involve separate regulatory pathways and evidence.

For a manufacturer, compliance is more than displaying certificates on a website. Review the product classification, quality-system scope, test reports, supplier controls, and production records. A factory making a plastic enclosure may face different controls from one assembling a sterile infusion pump. Details matter. Certification scope should match the actual site and product, and test results should be traceable to the relevant design or batch. Supplier documentation can be uneven, even when a company appears experienced. That deserves a closer look. Asking focused questions early can help buyers compare Chinese manufacturers on evidence, not promises, and identify which standards and approvals are genuinely relevant to their device.
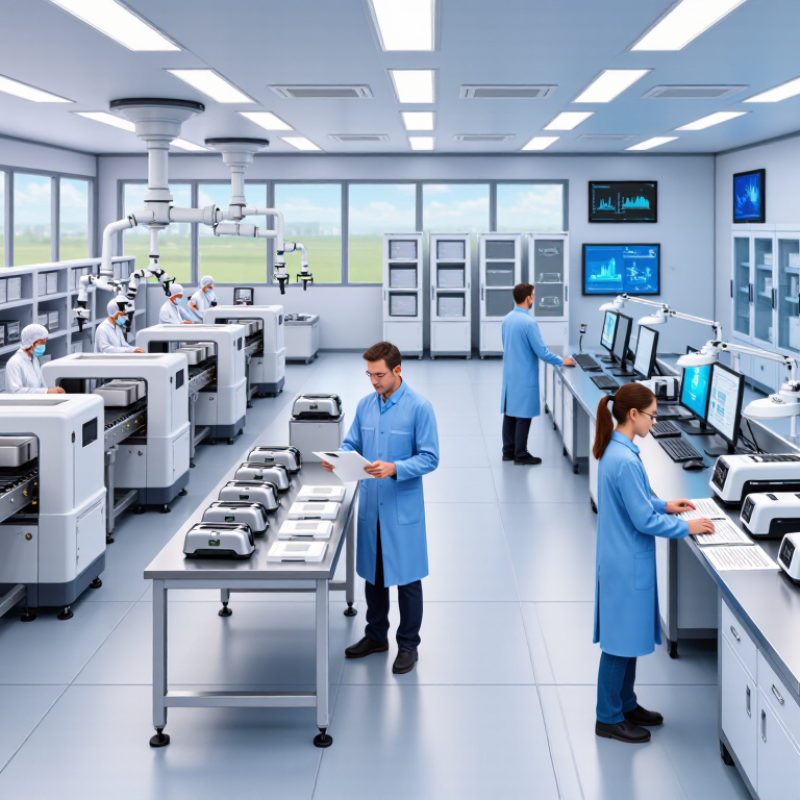
China Best Medical Manufacturing What Standards Apply?

Defining Medical Device Manufacturing in China

Defining Medical Device Manufacturing in China

Medical device manufacturing in China is more than assembling components. It covers controlled design transfer, supplier management, production, inspection, packaging, and records that connect each finished unit to its materials and process history.

WHO’s Medical Devices fact sheet describes roughly 2 million types of devices worldwide, grouped into more than 7,000 generic categories. That breadth helps explain why one checklist cannot fit every product.

In China, regulatory classification depends on intended use and risk. The applicable route may involve product registration or filing, alongside a manufacturing filing or license, depending on device class and circumstances. Manufacturers also need a quality system aligned with China’s medical-device Good Manufacturing Practice requirements.

ISO 13485 can support that system, while ISO 14971 provides a framework for risk management. Neither standard should be treated as a substitute for checking current NMPA requirements.

Standards and production controls

Standards are product-specific. Relevant national GB standards and industry YY standards may apply; some are mandatory, while others serve different roles. A practical review starts with the device’s intended purpose, materials, manufacturing steps, and claims. Then map those details to classification, testing, labeling, and production controls.

Small gaps matter. A supplier change, for example, can affect process validation or traceability. It is tempting to focus on certificates alone, but certificates do not prove every production batch is controlled.

WHO, Medical Devices fact sheet, provides the global device-category figures cited above.

How China Classifies Medical Devices by Risk

China classifies medical devices as Class I, II, or III according to risk and the level of control needed. Classification depends on intended use, body contact, duration of use, and operating mechanism. For example, reviewers may consider whether a device touches intact skin or enters the body, and how long it remains in contact. Class I generally presents lower risk; Class III includes higher-risk products, such as some implants. The product name alone does not determine its class.

Manufacturers should compare the device’s intended purpose and technical features with China’s current classification catalog and rules. Class I devices generally require filing, while Class II and III devices require registration. Exact requirements depend on the product and applicable guidance. Keep the classification rationale consistent with risk analysis, design records, and test evidence. Some cases are not straightforward; assumptions made too early can create delays.

Tips: Describe the intended use clearly. Record contact type and duration. Check the current catalog before planning submissions. If the classification remains unclear, seek guidance from the relevant authority. Small details matter.

China Medical Device Risk Classification: What Standards Apply?

China classifies medical devices into three classes according to risk to human health and the level of regulatory control required. Class I is the lowest-risk category, Class II is moderate risk, and Class III is the highest-risk category. The chart uses ordinal class rankings—not measured risk scores.

Which Regulations and Standards Apply to Manufacturers

Medical device manufacturers in China must match their obligations to the device’s intended use and risk classification. The National Medical Products Administration (NMPA) oversees registration or filing, while manufacturing sites must meet applicable quality management requirements. Class and product details matter. A change to materials, design, or intended use may affect the approval pathway, so teams should confirm requirements before production.

Manufacturers also need to identify applicable national GB standards and relevant industry YY standards. Some standards are mandatory; others may be recommended or apply only to particular products. Testing can cover safety, performance, biocompatibility, electrical risks, and packaging, depending on the device. ISO 13485 can support a quality management system, but it does not replace Chinese regulatory obligations. A checklist can still miss a detail; that is worth reviewing with qualified regulatory professionals.

Tips: Keep a standards register for each device, including standard numbers, editions, test evidence, and review dates. Check current NMPA notices and official standard publications before relying on older documents. Small details count. Labels, instructions, and manufacturing records should match the approved product information.

How Testing, Registration, and Certification Work

China Best Medical Manufacturing: What Standards Apply?

How Testing, Registration, and Certification Work

Testing, registration, and certification are related, but they are not interchangeable. A manufacturer may test a device against applicable standards, yet still need product registration or filing before market access. The route depends on the device’s intended use, risk class, and destination market. Small details matter: a change to a sensor, label, or sterilization method may require a review of existing evidence.

The World Health Organization’s Global Atlas of Medical Devices describes around two million different medical devices, grouped into more than 7,000 generic categories. That scale helps explain why regulators do not apply one checklist to every product. For a device made in China, teams commonly build a technical file with design records, risk analysis, test results, and manufacturing controls. ISO 13485 can support a quality management system, but certification alone does not approve a device for sale. Testing may cover electrical safety, biocompatibility, software, or sterilization, depending on the product.

Registration requirements also differ by market. China’s National Medical Products Administration uses risk-based classification and product-specific review pathways; other regions have their own rules and evidence expectations. Keep test reports traceable to the exact device version and intended use. That sounds obvious. In practice, document mismatches still happen, and they can delay review. A careful gap assessment before testing can prevent avoidable repeat work, though it cannot guarantee approval.

How to Assess a Manufacturer’s Compliance and Quality Systems

China Best Medical Manufacturing: What Standards Apply?
How to Assess a Manufacturer’s Compliance and Quality Systems

A certificate is a starting point, not a verdict. ISO’s 2022 Survey of Management System Standard Certifications counted more than 30,000 ISO 13485 certificates worldwide. That figure shows how widely the standard is used; it does not show how well a specific factory follows its procedures. Check that the certificate is current, covers the actual production site, and includes the device categories you plan to source. Then ask to see records, not just polished manuals. A quality system should connect staff training, supplier checks, calibration logs, production records, and corrective actions. Small gaps matter.

During an audit, trace one product lot from incoming materials to final release. Can the manufacturer identify the supplier, inspection results, equipment used, and person who approved the batch? Review complaint handling and corrective-action records for recurring problems and overdue investigations. WHO’s Global Patient Safety Report 2023 estimates that about one in ten patients is harmed in health care, underscoring why reliable production controls matter. This broad estimate is not a factory-performance measure. Still, it makes traceability and complaint review worth probing. A neat document set can hide weak habits on the production floor. Observe a shift change, ask operators how they report defects, and compare their answers with written procedures. Some audits miss the human details. That is a useful limitation to reflect on.

FAQS

What determines a medical device’s risk class in China?

Classification depends on intended use, body contact, contact duration, and how the device operates. The product name alone is not enough.

What are the three medical device classes?

Class I generally presents lower risk, while Class III includes higher-risk products, such as some implants. Class II sits between them.

Does each class follow the same market-entry route?

No. Class I devices generally require filing. Class II and III devices generally require registration, with exact requirements depending on the product.

What details should manufacturers record when assessing classification?

Record the intended purpose, contact type, and duration. Compare technical features with current classification rules. Small details matter.

Does passing a test mean a device is approved for sale?

No. Testing, registration, and certification serve different purposes. A device may still need product registration or filing.

What might device testing cover?

Testing may assess electrical safety, biocompatibility, software, or sterilization. The relevant tests depend on the product.

What should a quality certificate tell a buyer?

Check that it is current and covers the actual production site and relevant device categories. A certificate is only a starting point.

How can a buyer check factory traceability?

Follow one product lot from incoming materials to final release. Review supplier details, inspection results, equipment records, and batch approval.

What can an audit miss?

A tidy document set may not show daily habits. Watch a shift change and ask workers how they report defects. Audits have limits.

Conclusion

Medical device manufacturing in China involves more than assembling a product: manufacturers must manage design, production, quality control, and documentation in line with the device’s intended use and risk level. China classifies devices into three risk-based categories, with higher-risk products generally facing more extensive review and oversight. Understanding this classification is an essential first step in determining what standards apply to medical manufacturing, since obligations can vary by product type and production process.

Applicable requirements may include Chinese national or industry standards, quality management system controls, and relevant technical testing. Depending on the device, manufacturers may need to complete product testing, submit registration or filing materials, and obtain the appropriate approvals before placing it on the market. When assessing a manufacturer, review its quality system, production controls, traceability, testing records, and readiness to support regulatory submissions. A clear compliance process helps identify gaps early and supports consistent product quality throughout manufacturing.
Aria

Aria

Aria is a dedicated marketing professional with a deep passion for innovative strategies and a keen understanding of our company's product offerings. With a wealth of experience in the industry, Aria excels at crafting engaging content that highlights the unique features and benefits of our......